mutation predictions | marginal predictions | summary statistics | genome diff | command line log
|
breseq version 0.29.0
mutation predictions | marginal predictions | summary statistics | genome diff | command line log |
| read file | reads | bases | passed filters | average | longest | mapped | |
|---|---|---|---|---|---|---|---|
| errors | OtA1604_D37A0.5_R1_ | 6,044,593 | 604,459,300 | 100.0% | 100.0 bases | 100 bases | 64.5% |
| total | 6,044,593 | 604,459,300 | 100.0% | 100.0 bases | 100 bases | 64.5% |
| seq id | length | fit mean | fit dispersion | % mapped reads | description | ||
|---|---|---|---|---|---|---|---|
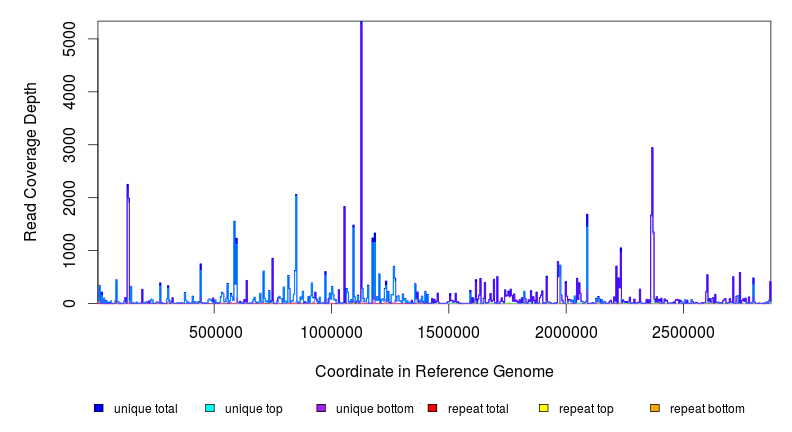
| coverage | distribution | NC_007793 | 2,872,769 | 76.1 | 221.7 | 99.8% | Staphylococcus aureus subsp. aureus USA300_FPR3757, complete |
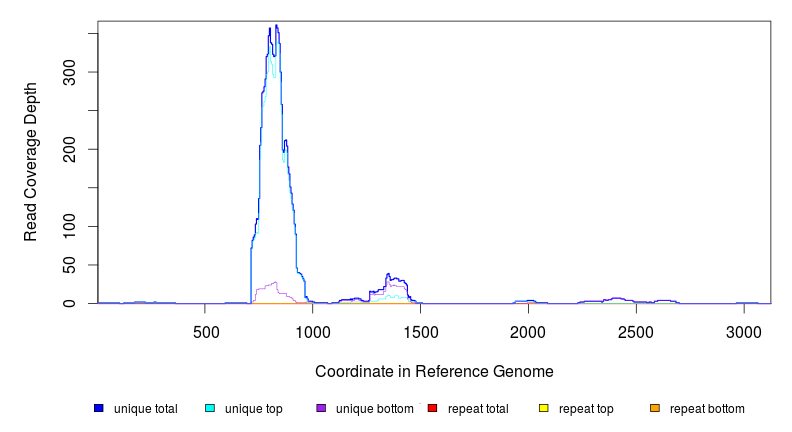
| coverage | distribution | NC_007790 | 3,125 | 2.2 | 1.9 | 0.0% | Staphylococcus aureus subsp. aureus USA300_FPR3757 plasmid pUSA01, |
| coverage | none aligned | NC_007791 | 4,439 | NA | NA | 0.0% | Staphylococcus aureus subsp. aureus USA300_FPR3757 plasmid pUSA02, |
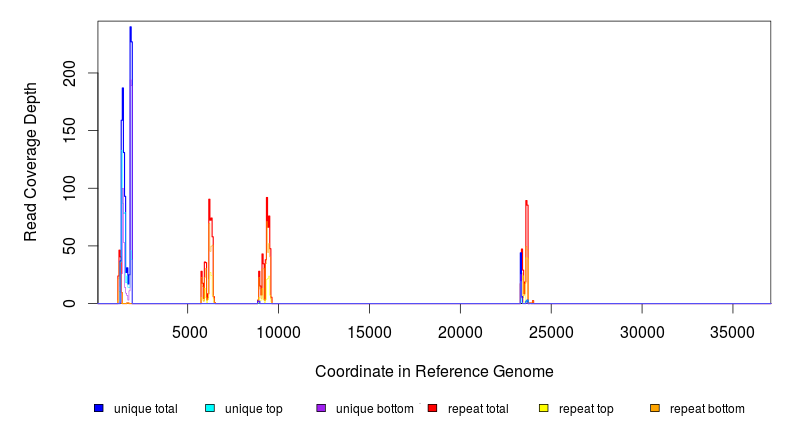
| coverage | distribution | NC_007792 | 37,136 | 30.1 | 2.6 | 0.0% | Staphylococcus aureus subsp. aureus USA300_FPR3757 plasmid pUSA03, |
| total | 2,917,469 | 100.0% |
fit dispersion is the ratio of the variance to the mean for the negative binomial fit. It is =1 for Poisson and >1 for over-dispersed data.
| option | value |
|---|---|
| Mode | Consensus/Mixed Base |
| Ploidy | 1 (haploid) |
| Consensus mutation E-value cutoff | 10 |
| Consensus frequency cutoff | 0.8 |
| Consensus minimum coverage each strand | OFF |
| Polymorphism E-value cutoff | 10 |
| Polymorphism frequency cutoff | 0.2 |
| Polymorphism minimum coverage each strand | OFF |
| Polymorphism bias cutoff | OFF |
| Predict indel polymorphisms | YES |
| Skip indel polymorphisms in homopolymers runs of | OFF |
| Skip base substitutions when they create a homopolymer flanked on each side by | OFF |
| step | start | end | elapsed |
|---|---|---|---|
| Read and reference sequence file input | 15:35:31 03 Mar 2017 | 15:36:27 03 Mar 2017 | 56 seconds |
| Read alignment to reference genome | 15:36:28 03 Mar 2017 | 15:47:50 03 Mar 2017 | 11 minutes 22 seconds |
| Resolving alignments with junction candidates | 15:47:50 03 Mar 2017 | 15:49:41 03 Mar 2017 | 1 minute 51 seconds |
| Creating BAM files | 15:49:41 03 Mar 2017 | 15:51:01 03 Mar 2017 | 1 minute 20 seconds |
| Tabulating error counts | 15:51:01 03 Mar 2017 | 15:51:49 03 Mar 2017 | 48 seconds |
| Re-calibrating base error rates | 15:51:49 03 Mar 2017 | 15:51:52 03 Mar 2017 | 3 seconds |
| Examining read alignment evidence | 15:51:52 03 Mar 2017 | 16:00:32 03 Mar 2017 | 8 minutes 40 seconds |
| Polymorphism statistics | 16:00:32 03 Mar 2017 | 16:00:38 03 Mar 2017 | 6 seconds |
| Output | 16:00:38 03 Mar 2017 | 16:13:09 03 Mar 2017 | 12 minutes 31 seconds |
| Total | 37 minutes 37 seconds | ||