mutation predictions | marginal predictions | summary statistics | genome diff | command line log
|
breseq version 0.35.4
mutation predictions | marginal predictions | summary statistics | genome diff | command line log |
| read file | reads | bases | passed filters | average | longest | mapped | |
|---|---|---|---|---|---|---|---|
| errors | ATCC_S10_L001_R1_001 | 902,159 | 194,754,060 | 99.9% | 215.9 bases | 251 bases | 98.2% |
| total | 902,159 | 194,754,060 | 99.9% | 215.9 bases | 251 bases | 98.2% |
| seq id | length | fit mean | fit dispersion | % mapped reads | description | ||
|---|---|---|---|---|---|---|---|
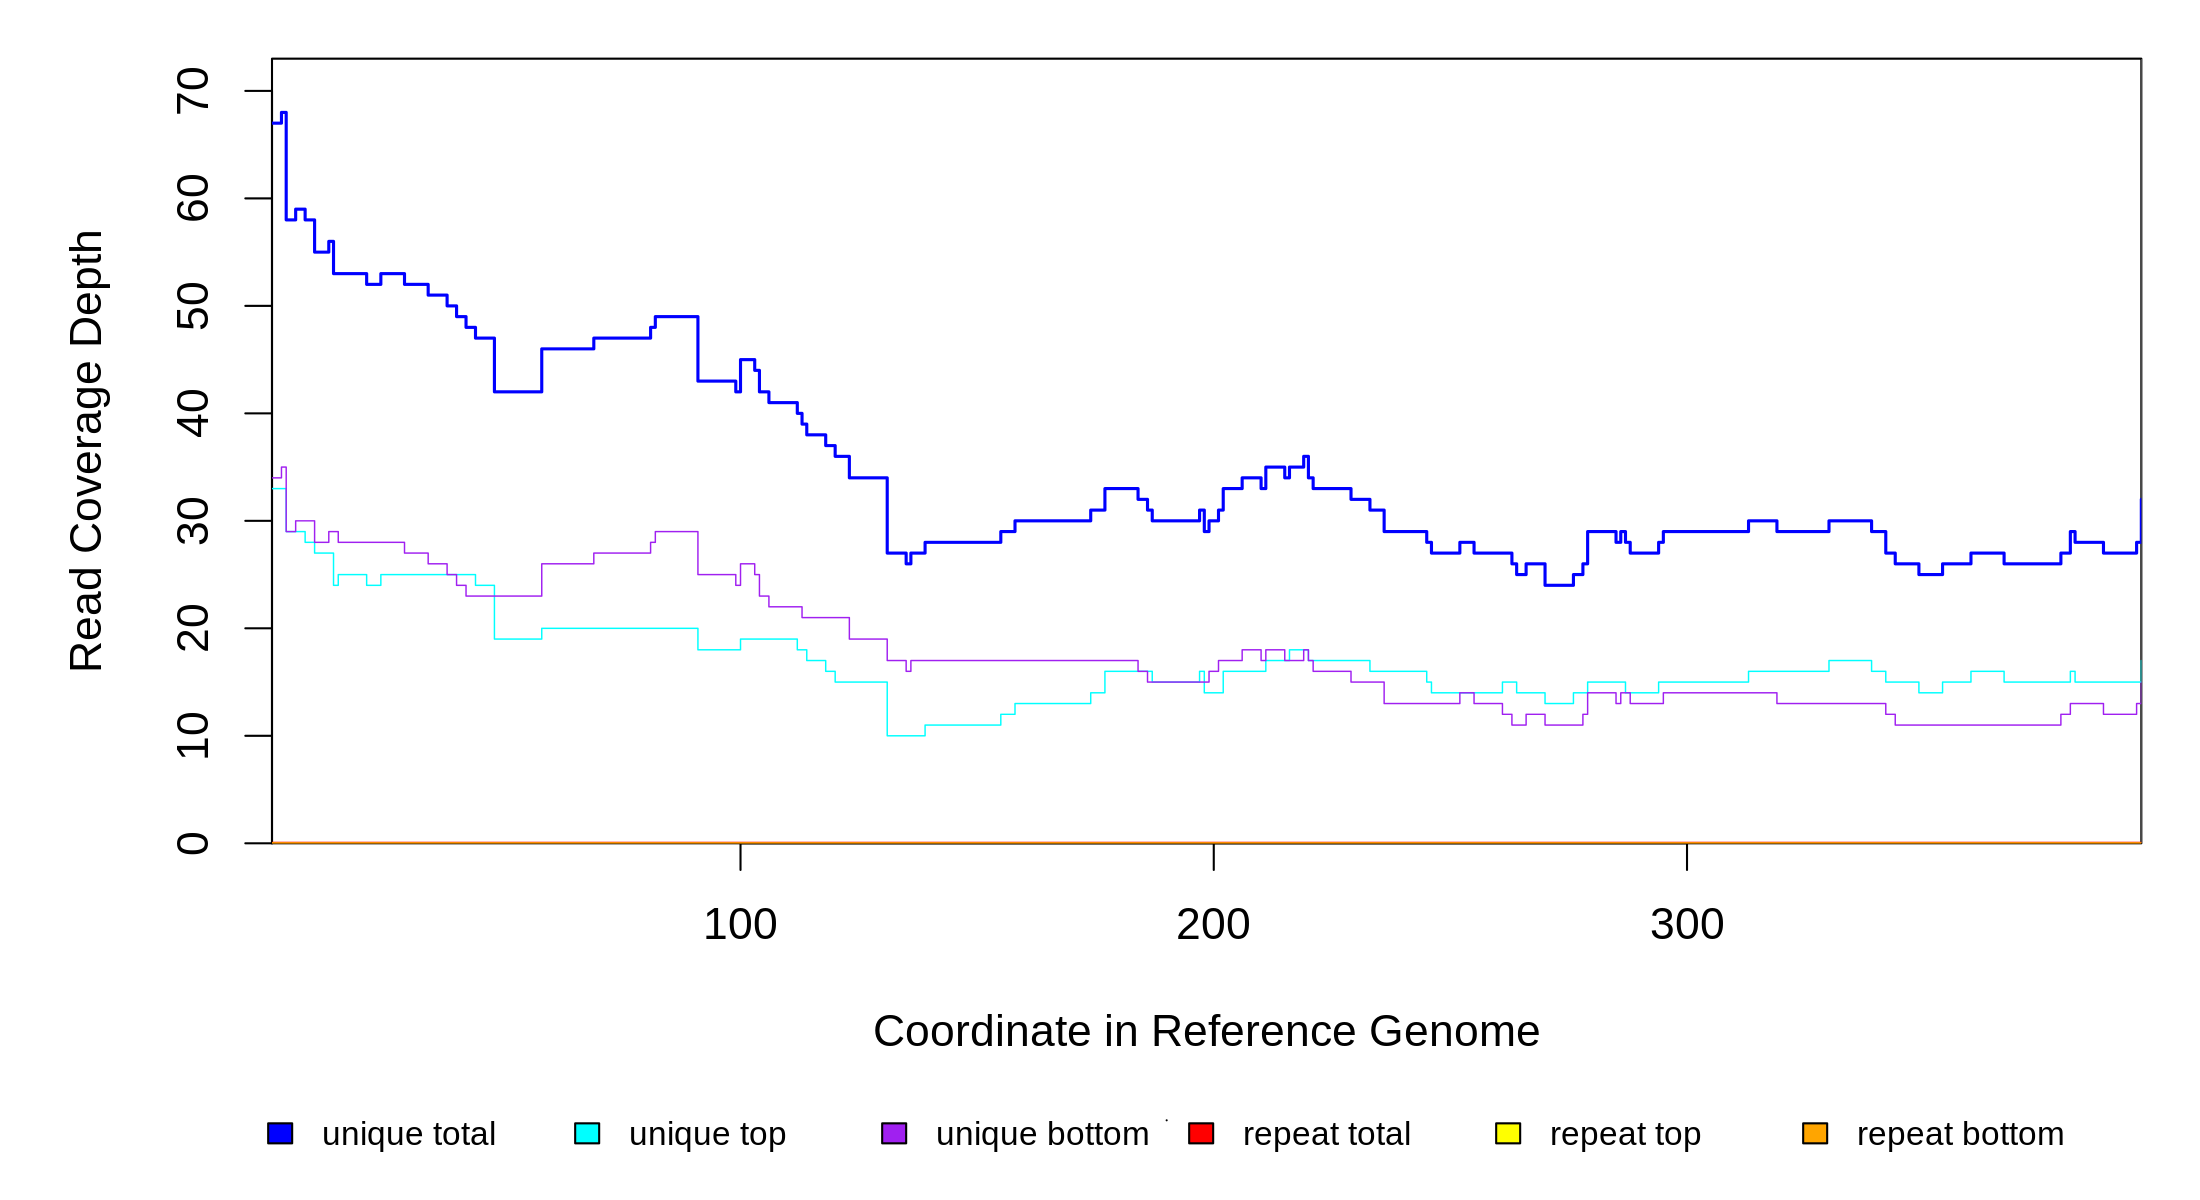
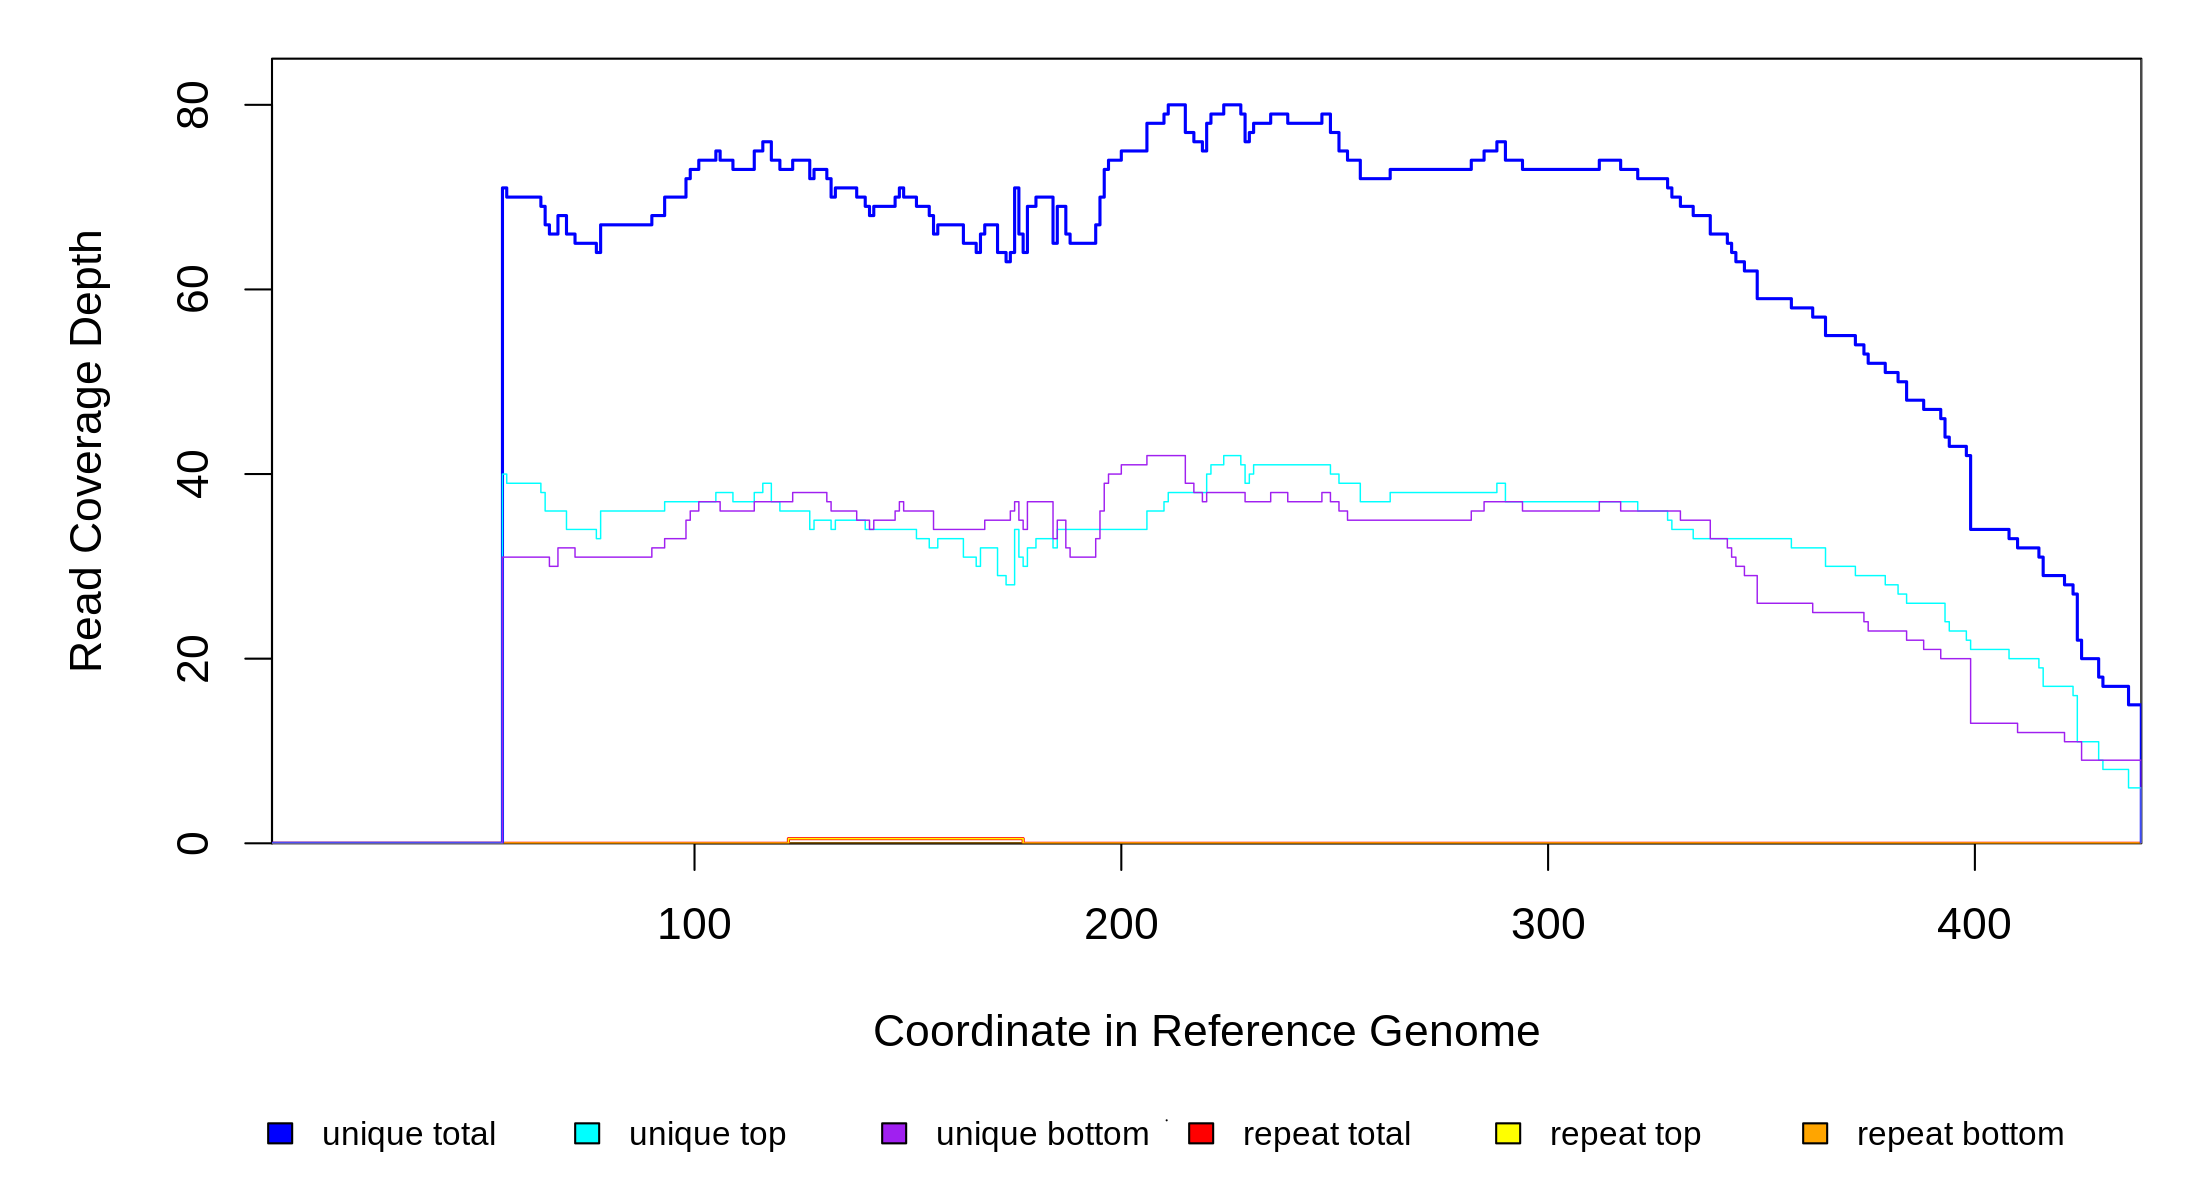
| coverage | distribution | 6 | 396 | [34.7] | [2.6] | 0.0% | Aggregatibacter actinomycetemcomitans Aa23R |
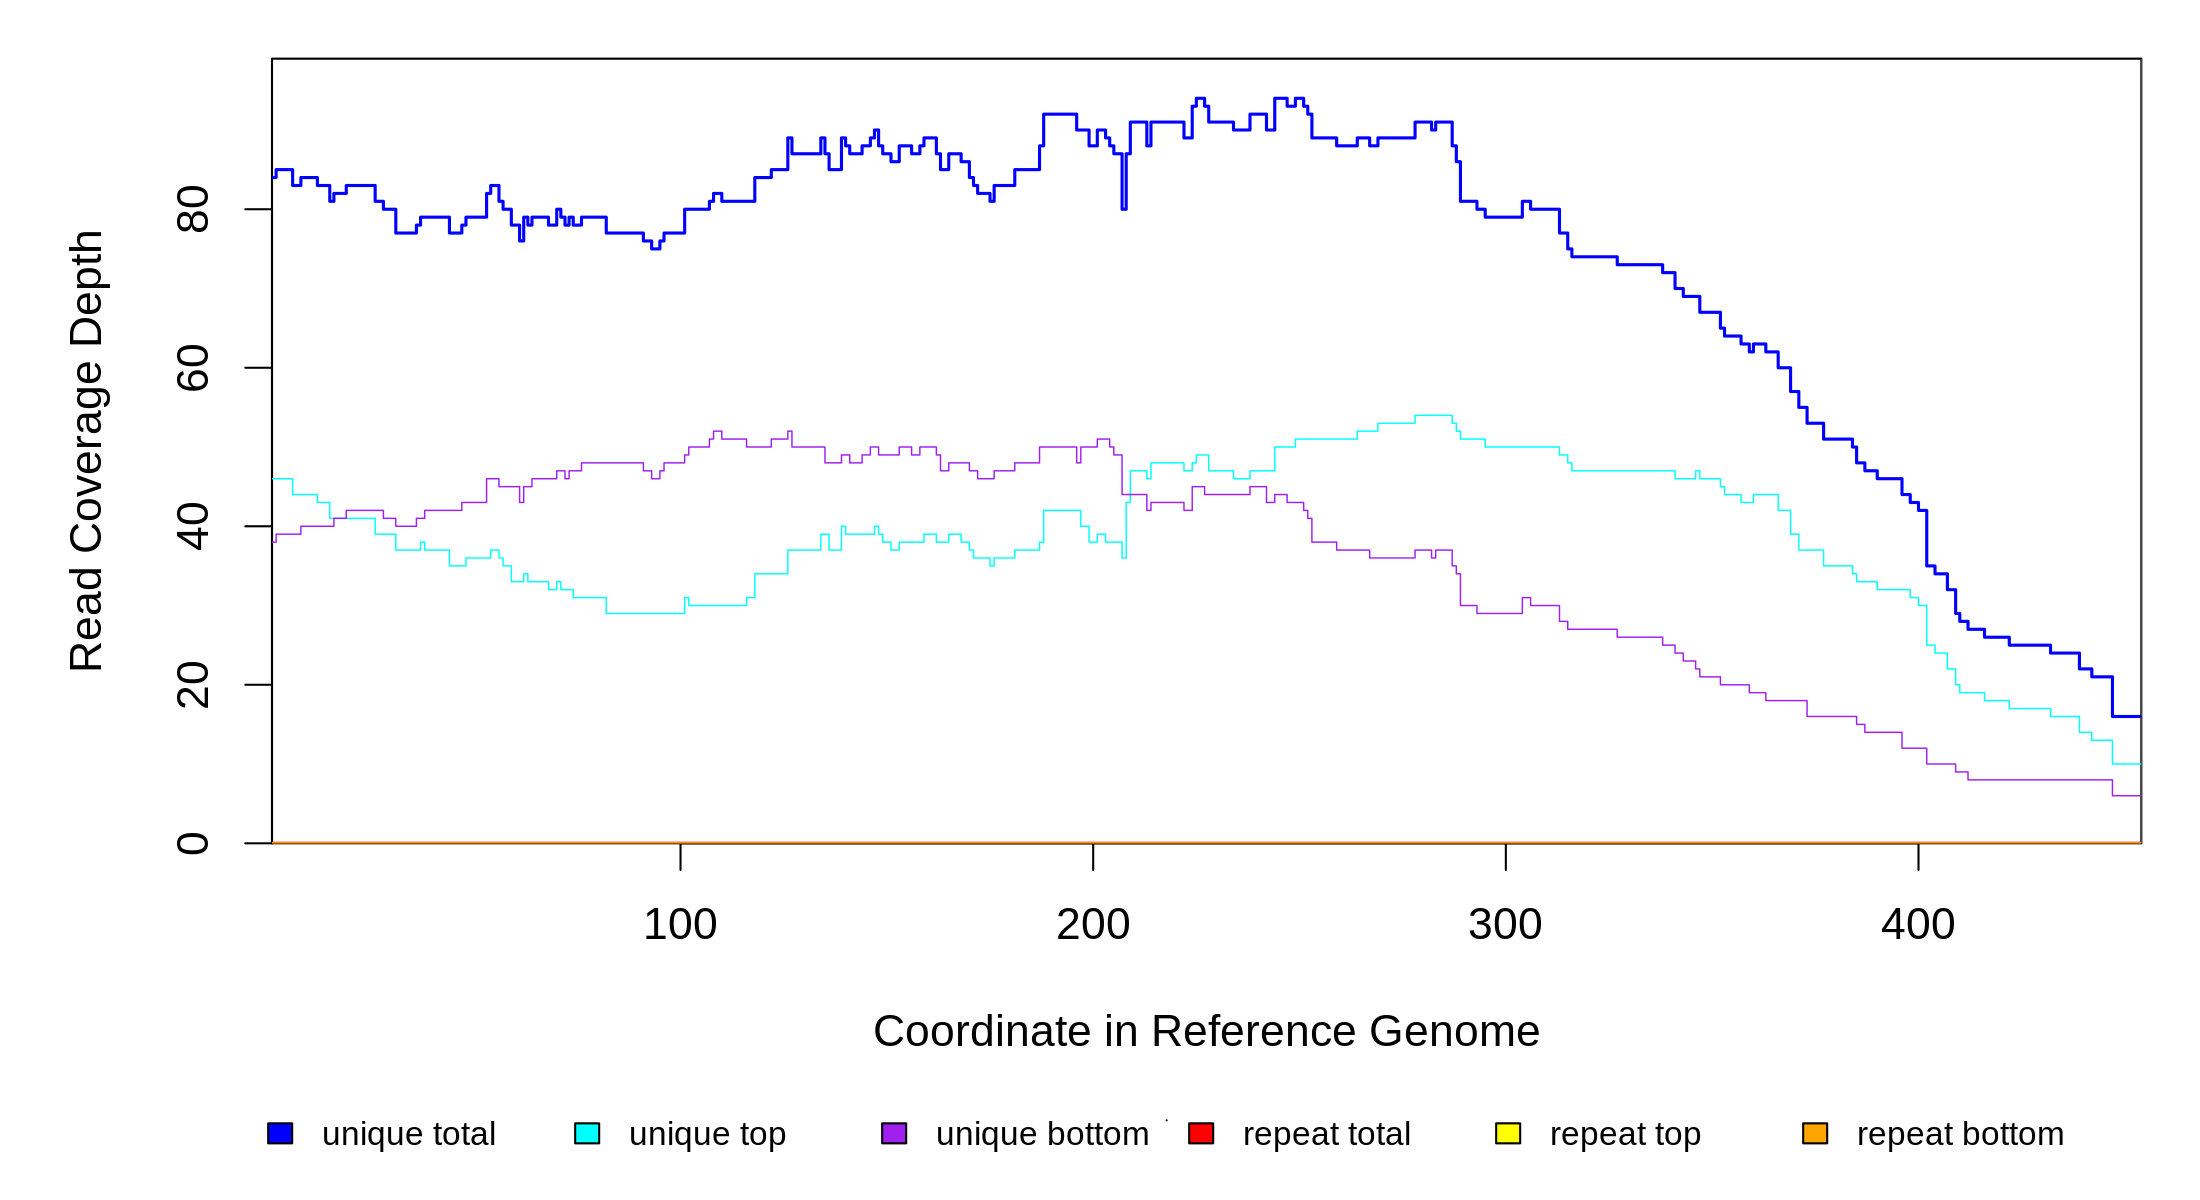
| coverage | distribution | 4 | 454 | 84.9 | 1.0 | 0.0% | Aggregatibacter actinomycetemcomitans Aa23R |
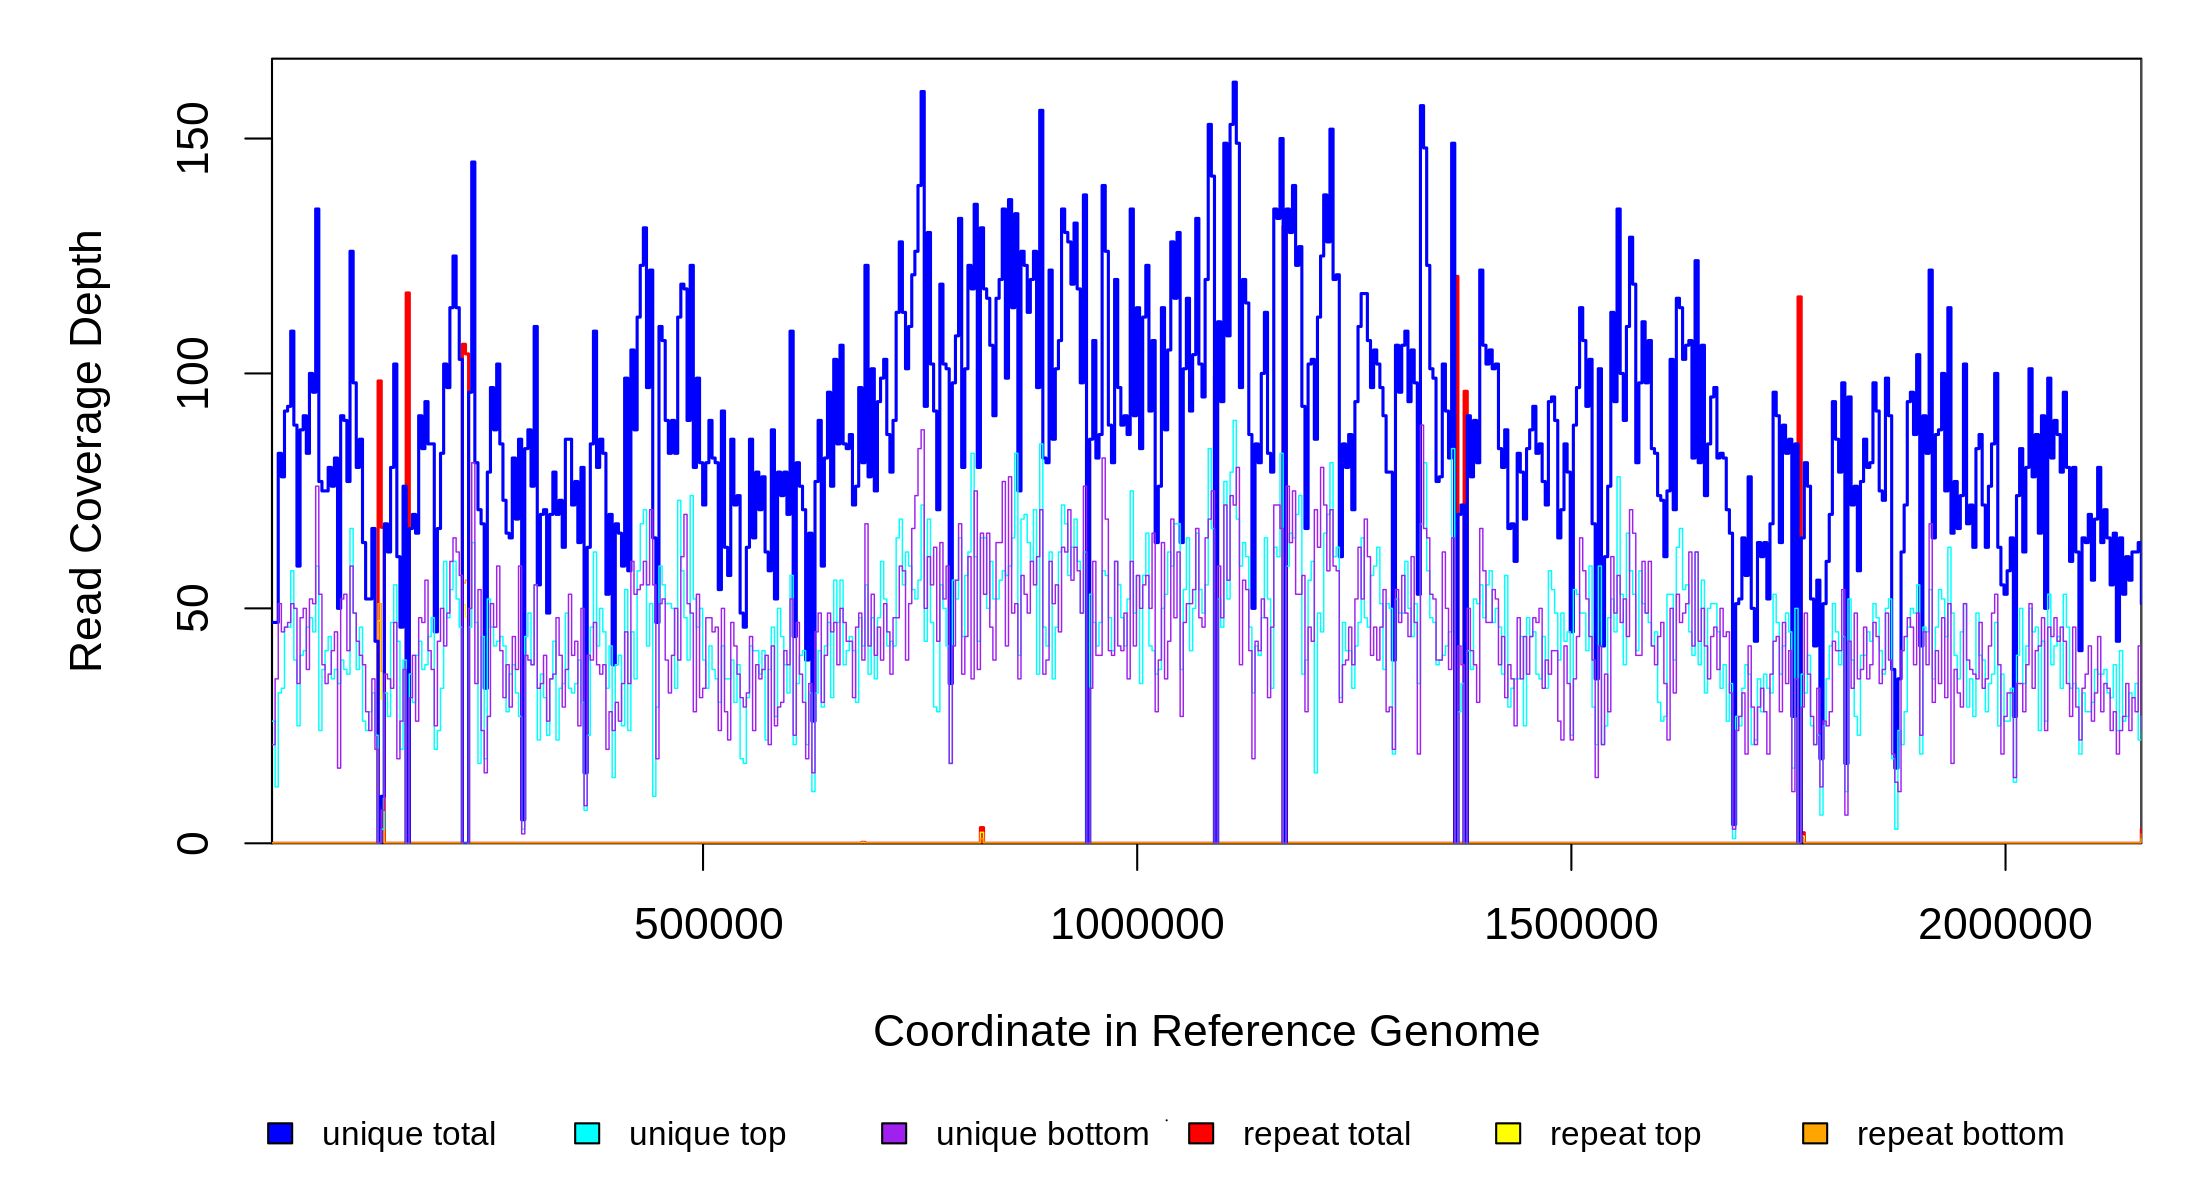
| coverage | distribution | 1 | 2,156,994 | 90.9 | 6.5 | 100.0% | Aggregatibacter actinomycetemcomitans Aa23R |
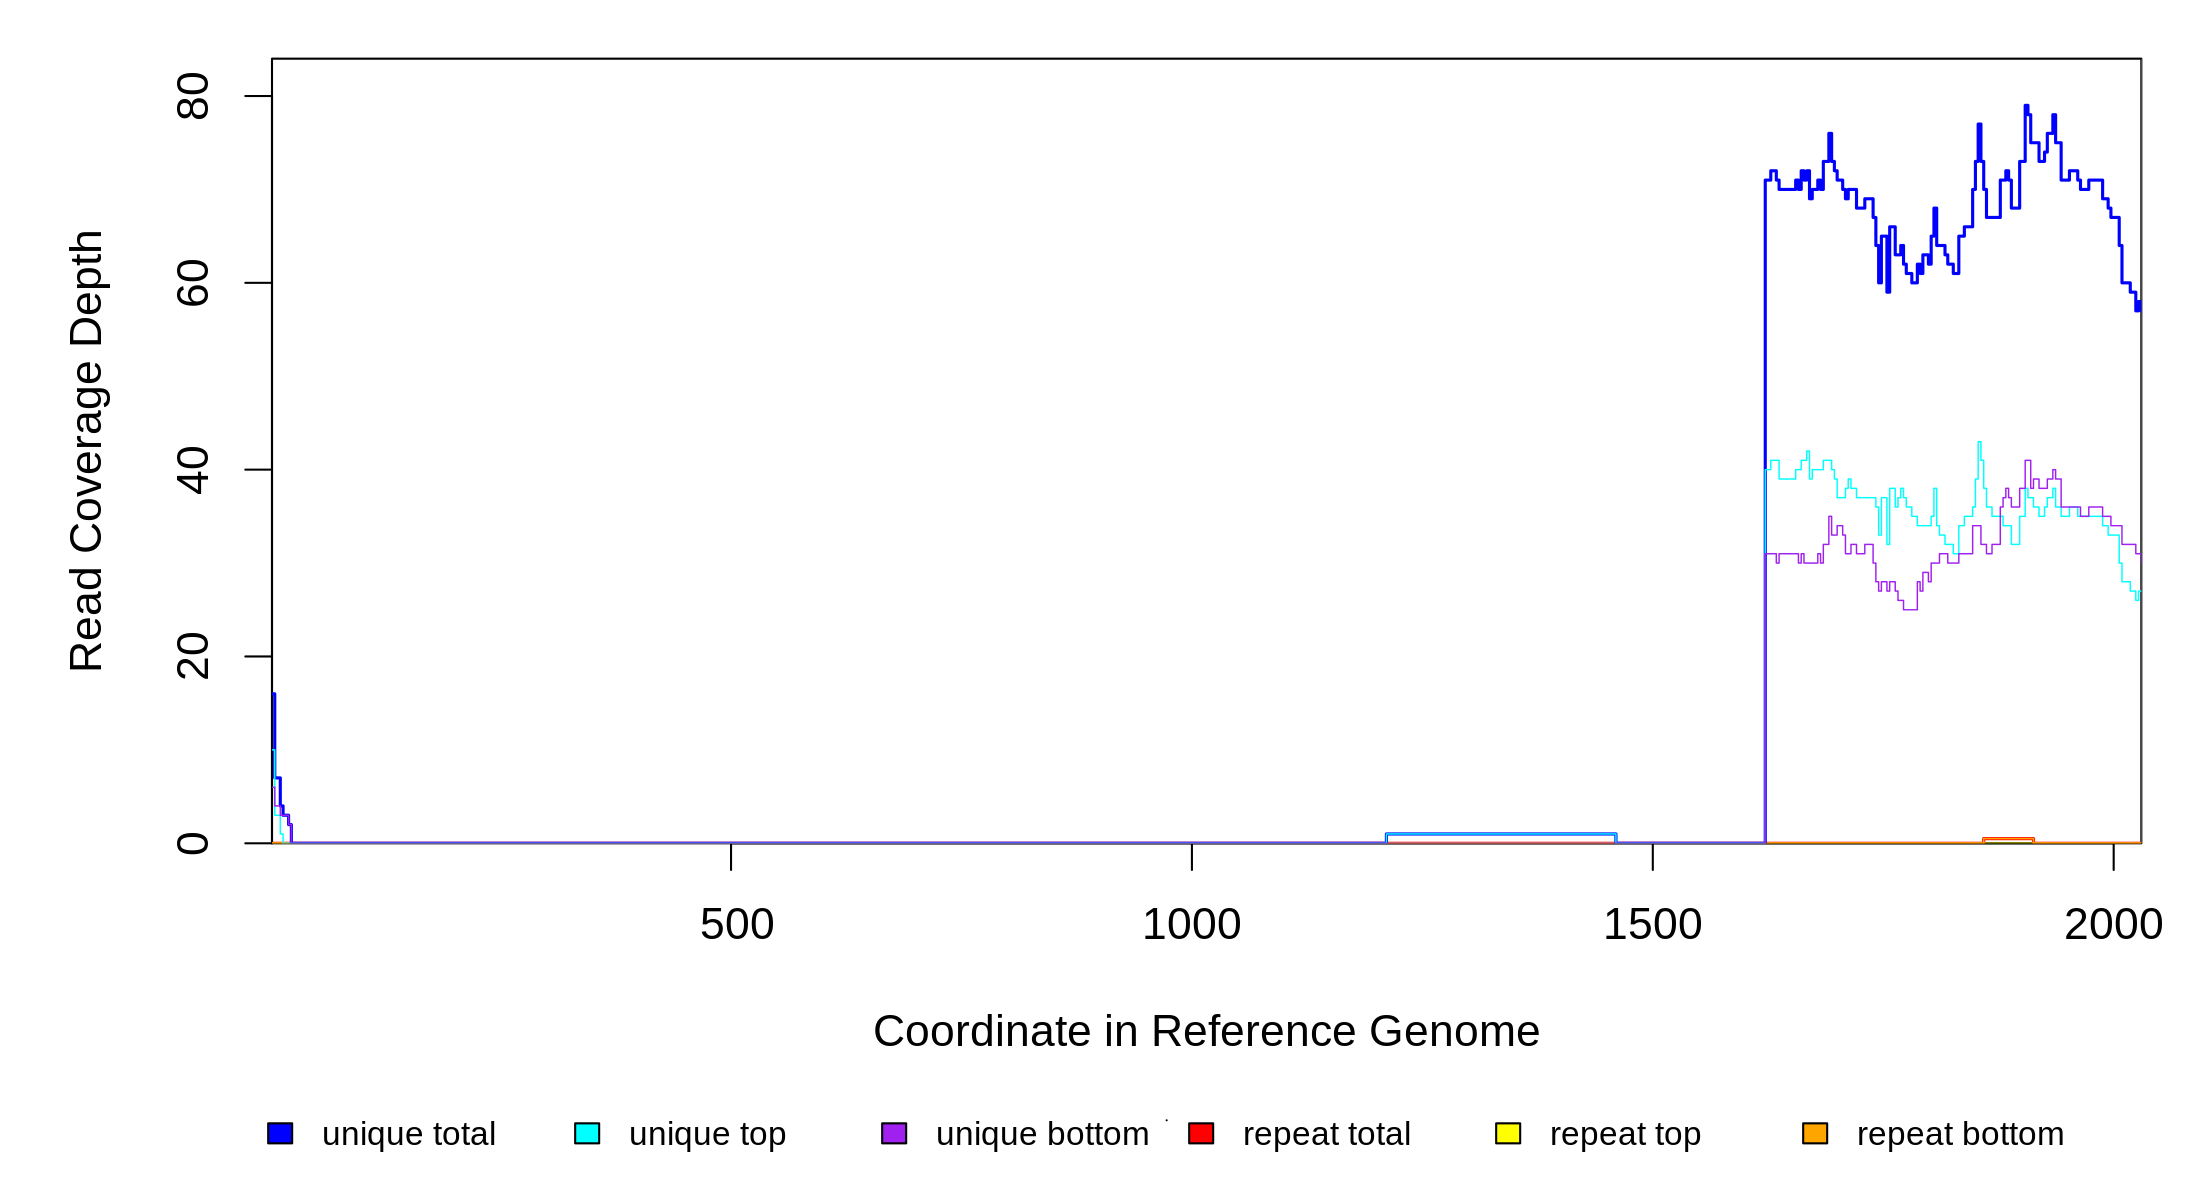
| coverage | distribution | 3 | 2,032 | 69.4 | 1.0 | 0.0% | Aggregatibacter actinomycetemcomitans Aa23R |
| coverage | distribution | 2 | 4,576 | NA | NA | 0.0% | Aggregatibacter actinomycetemcomitans Aa23R |
| coverage | distribution | 5 | 439 | 74.1 | 1.0 | 0.0% | Aggregatibacter actinomycetemcomitans Aa23R |
| total | 2,164,891 | 100.0% |
fit dispersion is the ratio of the variance to the mean for the negative binomial fit. It is =1 for Poisson and >1 for over-dispersed data.
Fit failed Negative binomial fit failed for this reference sequence. It may have an unusual coverage depth distribution. JC and MC predictions may be less accurate.
Insufficient coverage Reference sequence counted as entirely deleted due to low coverage. Try either the -t,--targeted-sequencing or the -c,--contig-reference option if you want mutations called for these reference sequences.
| option | limit | actual |
|---|---|---|
| Number of alignment pairs examined for constructing junction candidates | ≤ 100000 | 12565 |
| Coverage evenness (position-hash) score of junction candidates | ≥ 2 | ≥ 2 |
| Test this many junction candidates (n). May be smaller if not enough passed the coverage evenness threshold | 100 ≤ n ≤ 5000 | 297 |
| Total length of all junction candidates (factor times the reference genome length) | ≤ 0.1 | 0.070 |
| reference sequence | pr(no read start) |
|---|---|
| 6 | NA |
| 4 | 0.92181 |
| 1 | 0.85882 |
| 3 | 0.98786 |
| 2 | NA |
| 5 | 0.94661 |
pr(no read start) is the probability that there will not be an aligned read whose first base matches a given position on a given strand.
| option | value |
|---|---|
| Coverage evenness (position-hash) score of predicted junctions must be | ≥ 3 |
| Minimum probablilty assigned that no mapped read will start at a given position and strand for junction prediction | 0.1 |
| Junction allow suboptimal matches | FALSE |
| Skew score of predicted junction (−log10 probability of unusual coverage evenness) must be | ≤ 3 |
| Number of bases that at least one read must overlap each uniquely aligned side of a predicted junction | ≥ 1 |
| option | value |
|---|---|
| Mode | Consensus/Mixed Base |
| Ploidy | 1 (haploid) |
| Consensus mutation E-value cutoff | 10 |
| Consensus frequency cutoff | 0.8 |
| Consensus minimum variant coverage each strand | OFF |
| Consensus minimum total coverage each strand | OFF |
| Consensus minimum variant coverage | OFF |
| Consensus minimum total coverage | OFF |
| Polymorphism E-value cutoff | 10 |
| Polymorphism frequency cutoff | 0.2 |
| Polymorphism minimum variant coverage each strand | OFF |
| Polymorphism minimum total coverage each strand | OFF |
| Polymorphism minimum variant coverage | OFF |
| Polymorphism minimum total coverage | OFF |
| Polymorphism bias cutoff | OFF |
| Predict indel polymorphisms | YES |
| Skip indel polymorphisms in homopolymers runs of | OFF |
| Skip base substitutions when they create a homopolymer flanked on each side by | OFF |
| program | version |
|---|---|
| bowtie2 | 2.4.1 |
| R | 4.0.0 |
| step | start | end | elapsed |
|---|---|---|---|
| Read and reference sequence file input | 10:20:22 07 Feb 2022 | 10:20:41 07 Feb 2022 | 19 seconds |
| Read alignment to reference genome | 10:20:41 07 Feb 2022 | 10:21:36 07 Feb 2022 | 55 seconds |
| Preprocessing alignments for candidate junction identification | 10:21:36 07 Feb 2022 | 10:21:54 07 Feb 2022 | 18 seconds |
| Preliminary analysis of coverage distribution | 10:21:54 07 Feb 2022 | 10:22:45 07 Feb 2022 | 51 seconds |
| Identifying junction candidates | 10:22:45 07 Feb 2022 | 10:22:54 07 Feb 2022 | 9 seconds |
| Re-alignment to junction candidates | 10:22:54 07 Feb 2022 | 10:23:16 07 Feb 2022 | 22 seconds |
| Resolving best read alignments | 10:23:16 07 Feb 2022 | 10:23:42 07 Feb 2022 | 26 seconds |
| Creating BAM files | 10:23:42 07 Feb 2022 | 10:24:20 07 Feb 2022 | 38 seconds |
| Tabulating error counts | 10:24:20 07 Feb 2022 | 10:24:40 07 Feb 2022 | 20 seconds |
| Re-calibrating base error rates | 10:24:40 07 Feb 2022 | 10:24:47 07 Feb 2022 | 7 seconds |
| Examining read alignment evidence | 10:24:47 07 Feb 2022 | 10:28:40 07 Feb 2022 | 3 minutes 53 seconds |
| Polymorphism statistics | 10:28:40 07 Feb 2022 | 10:28:41 07 Feb 2022 | 1 second |
| Output | 10:28:41 07 Feb 2022 | 10:28:59 07 Feb 2022 | 18 seconds |
| Total | 8 minutes 37 seconds | ||