mutation predictions | marginal predictions | summary statistics | genome diff | command line log
|
breseq version 0.35.4
mutation predictions | marginal predictions | summary statistics | genome diff | command line log |
| read file | reads | bases | passed filters | average | longest | mapped | |
|---|---|---|---|---|---|---|---|
| errors | ATCC_S10_L001_R1_001 | 902,159 | 194,754,060 | 99.9% | 215.9 bases | 251 bases | 93.2% |
| total | 902,159 | 194,754,060 | 99.9% | 215.9 bases | 251 bases | 93.2% |
| seq id | length | fit mean | fit dispersion | % mapped reads | description | ||
|---|---|---|---|---|---|---|---|
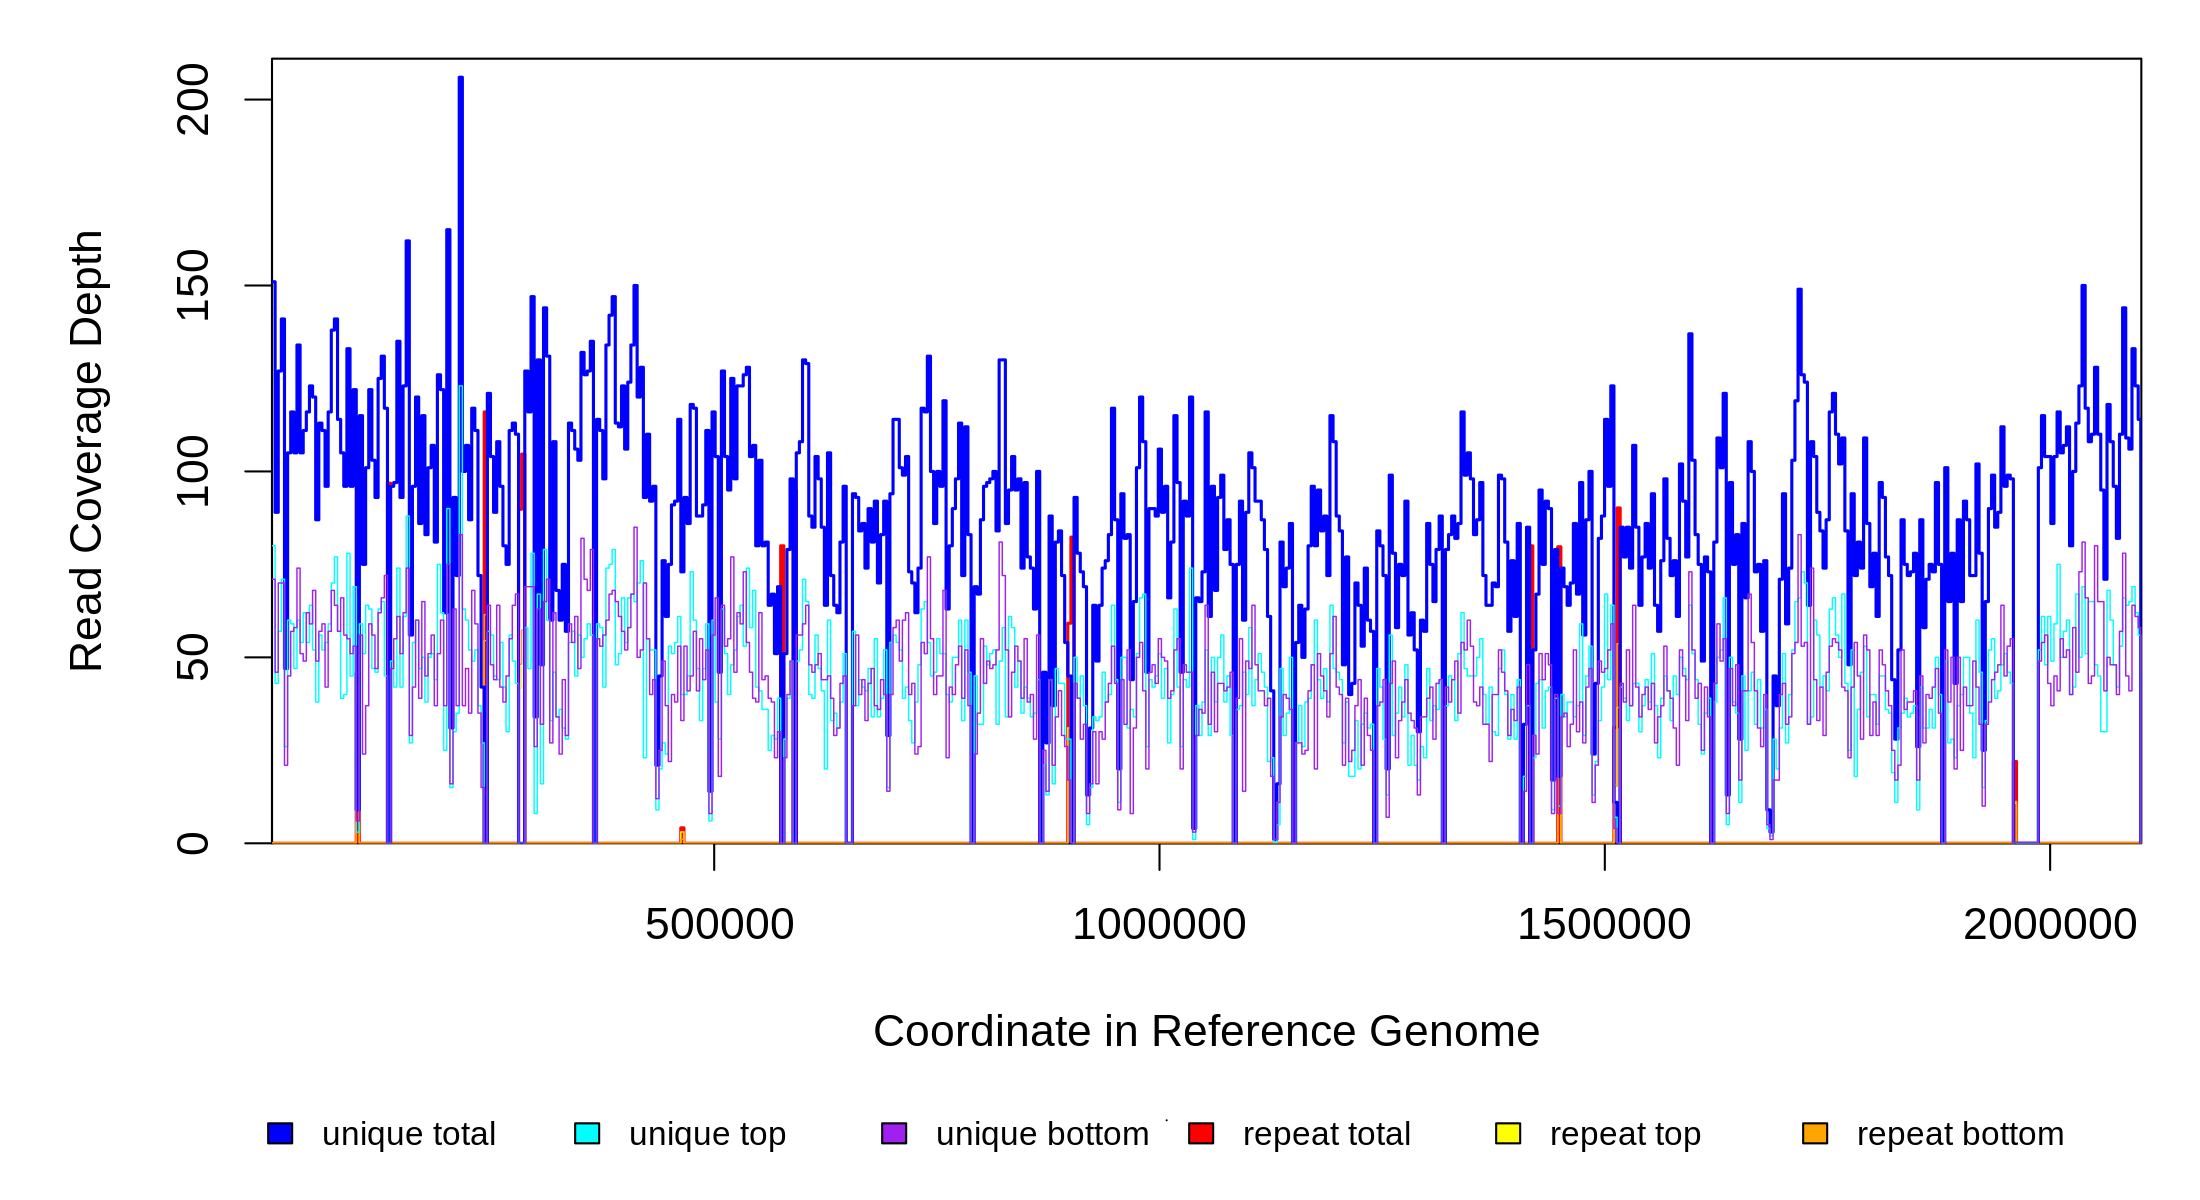
| coverage | distribution | 1 | 2,102,524 | 91.3 | 6.2 | 100.0% | Aggregatibacter actinomycetemcomitans Aa31S |
| coverage | distribution | 2 | 196 | NA | NA | 0.0% | Aggregatibacter actinomycetemcomitans Aa31S |
| total | 2,102,720 | 100.0% |
fit dispersion is the ratio of the variance to the mean for the negative binomial fit. It is =1 for Poisson and >1 for over-dispersed data.
Insufficient coverage Reference sequence counted as entirely deleted due to low coverage. Try either the -t,--targeted-sequencing or the -c,--contig-reference option if you want mutations called for these reference sequences.
| option | limit | actual |
|---|---|---|
| Number of alignment pairs examined for constructing junction candidates | ≤ 100000 | 17168 |
| Coverage evenness (position-hash) score of junction candidates | ≥ 2 | ≥ 2 |
| Test this many junction candidates (n). May be smaller if not enough passed the coverage evenness threshold | 100 ≤ n ≤ 5000 | 371 |
| Total length of all junction candidates (factor times the reference genome length) | ≤ 0.1 | 0.090 |
| reference sequence | pr(no read start) |
|---|---|
| 1 | 0.86331 |
| 2 | NA |
pr(no read start) is the probability that there will not be an aligned read whose first base matches a given position on a given strand.
| option | value |
|---|---|
| Coverage evenness (position-hash) score of predicted junctions must be | ≥ 3 |
| Minimum probablilty assigned that no mapped read will start at a given position and strand for junction prediction | 0.1 |
| Junction allow suboptimal matches | FALSE |
| Skew score of predicted junction (−log10 probability of unusual coverage evenness) must be | ≤ 3 |
| Number of bases that at least one read must overlap each uniquely aligned side of a predicted junction | ≥ 1 |
| option | value |
|---|---|
| Mode | Consensus/Mixed Base |
| Ploidy | 1 (haploid) |
| Consensus mutation E-value cutoff | 10 |
| Consensus frequency cutoff | 0.8 |
| Consensus minimum variant coverage each strand | OFF |
| Consensus minimum total coverage each strand | OFF |
| Consensus minimum variant coverage | OFF |
| Consensus minimum total coverage | OFF |
| Polymorphism E-value cutoff | 10 |
| Polymorphism frequency cutoff | 0.2 |
| Polymorphism minimum variant coverage each strand | OFF |
| Polymorphism minimum total coverage each strand | OFF |
| Polymorphism minimum variant coverage | OFF |
| Polymorphism minimum total coverage | OFF |
| Polymorphism bias cutoff | OFF |
| Predict indel polymorphisms | YES |
| Skip indel polymorphisms in homopolymers runs of | OFF |
| Skip base substitutions when they create a homopolymer flanked on each side by | OFF |
| program | version |
|---|---|
| bowtie2 | 2.4.1 |
| R | 4.0.0 |
| step | start | end | elapsed |
|---|---|---|---|
| Read and reference sequence file input | 14:43:24 20 Jan 2022 | 14:43:42 20 Jan 2022 | 18 seconds |
| Read alignment to reference genome | 14:43:43 20 Jan 2022 | 14:44:34 20 Jan 2022 | 51 seconds |
| Preprocessing alignments for candidate junction identification | 14:44:34 20 Jan 2022 | 14:44:51 20 Jan 2022 | 17 seconds |
| Preliminary analysis of coverage distribution | 14:44:51 20 Jan 2022 | 14:45:34 20 Jan 2022 | 43 seconds |
| Identifying junction candidates | 14:45:34 20 Jan 2022 | 14:46:00 20 Jan 2022 | 26 seconds |
| Re-alignment to junction candidates | 14:46:00 20 Jan 2022 | 14:46:18 20 Jan 2022 | 18 seconds |
| Resolving best read alignments | 14:46:18 20 Jan 2022 | 14:46:44 20 Jan 2022 | 26 seconds |
| Creating BAM files | 14:46:44 20 Jan 2022 | 14:47:20 20 Jan 2022 | 36 seconds |
| Tabulating error counts | 14:47:20 20 Jan 2022 | 14:47:38 20 Jan 2022 | 18 seconds |
| Re-calibrating base error rates | 14:47:38 20 Jan 2022 | 14:47:39 20 Jan 2022 | 1 second |
| Examining read alignment evidence | 14:47:39 20 Jan 2022 | 14:51:15 20 Jan 2022 | 3 minutes 36 seconds |
| Polymorphism statistics | 14:51:15 20 Jan 2022 | 14:51:16 20 Jan 2022 | 1 second |
| Output | 14:51:16 20 Jan 2022 | 14:52:07 20 Jan 2022 | 51 seconds |
| Total | 8 minutes 42 seconds | ||