mutation predictions | marginal predictions | summary statistics | genome diff | command line log
|
breseq version 0.38.1
mutation predictions | marginal predictions | summary statistics | genome diff | command line log |
| read file | reads | bases | passed filters | average | longest | mapped | |
|---|---|---|---|---|---|---|---|
| errors | Fusobacterium-ATCC-fusnuc_S12_L001_R1_001 | 918,623 | 203,508,046 | 100.0% | 221.5 bases | 251 bases | 99.2% |
| errors | Fusobacterium-ATCC-fusnuc_S12_L001_R2_001 | 918,623 | 203,451,002 | 100.0% | 221.5 bases | 251 bases | 99.1% |
| total | 1,837,246 | 406,959,048 | 100.0% | 221.5 bases | 251 bases | 99.2% |
| seq id | length | fit mean | fit relative_variance | % mapped reads | description | ||
|---|---|---|---|---|---|---|---|
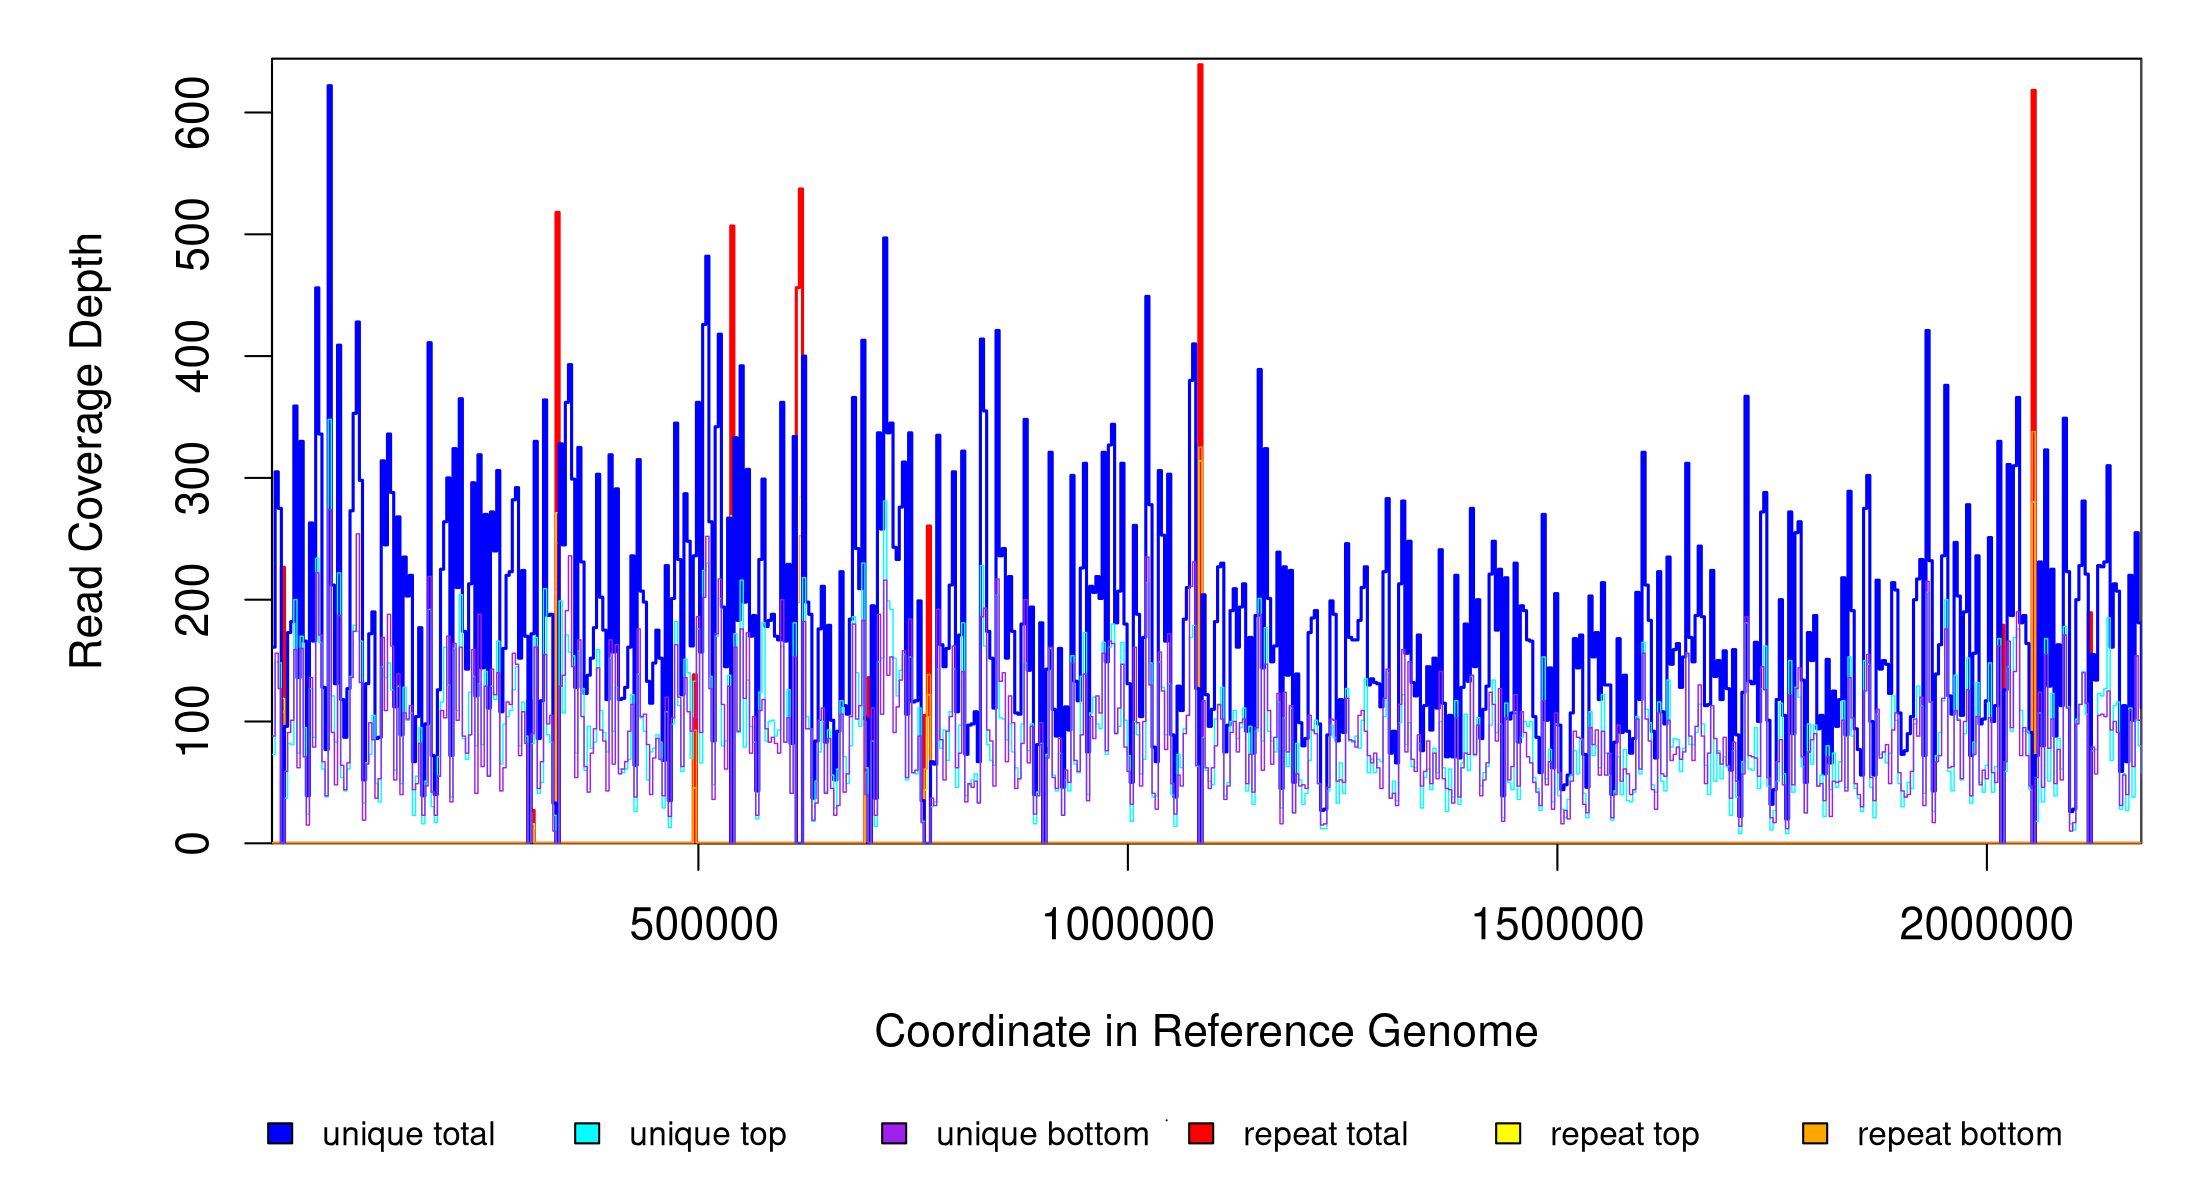
| coverage | distribution | CP028101 | 2,180,101 | 177.5 | 47.5 | 100.0% | Fusobacterium nucleatum subsp. nucleatum ATCC 25586 chromosome, |
| total | 2,180,101 | 100.0% |
fit relative_variance is the ratio of the variance to the mean for the negative binomial fit. It is =1 for Poisson and >1 for over-dispersed data.
| option | limit | actual |
|---|---|---|
| Number of alignment pairs examined for constructing junction candidates | ≤ 100000 | 4226 |
| Coverage evenness (position-hash) score of junction candidates | ≥ 2 | ≥ 3 |
| Test this many junction candidates (n). May be smaller if not enough passed the coverage evenness threshold | 100 ≤ n ≤ 5000 | 8 |
| Total length of all junction candidates (factor times the reference genome length) | ≤ 0.1 | 0.002 |
| reference sequence | pr(no read start) |
|---|---|
| CP028101 | 0.78510 |
pr(no read start) is the probability that there will not be an aligned read whose first base matches a given position on a given strand.
| option | value |
|---|---|
| Coverage evenness (position-hash) score of predicted junctions must be | ≥ 3 |
| Minimum probablilty assigned that no mapped read will start at a given position and strand for junction prediction | 0.1 |
| Junction allow suboptimal matches | FALSE |
| Skew score of predicted junction (−log10 probability of unusual coverage evenness) must be | ≤ 3 |
| Number of bases that at least one read must overlap each uniquely aligned side of a predicted junction | ≥ 1 |
| option | value |
|---|---|
| Mode | Consensus/Mixed Base |
| Ploidy | 1 (haploid) |
| Consensus mutation E-value cutoff | 10 |
| Consensus frequency cutoff | 0.8 |
| Consensus minimum variant coverage each strand | OFF |
| Consensus minimum total coverage each strand | OFF |
| Consensus minimum variant coverage | OFF |
| Consensus minimum total coverage | OFF |
| Polymorphism E-value cutoff | 10 |
| Polymorphism frequency cutoff | 0.2 |
| Polymorphism minimum variant coverage each strand | OFF |
| Polymorphism minimum total coverage each strand | OFF |
| Polymorphism minimum variant coverage | OFF |
| Polymorphism minimum total coverage | OFF |
| Polymorphism bias cutoff | OFF |
| Predict indel polymorphisms | YES |
| Skip indel polymorphisms in homopolymers runs of | OFF |
| Skip base substitutions when they create a homopolymer flanked on each side by | OFF |
| program | version |
|---|---|
| bowtie2 | s.2023.01/rocky8/x86_64/amd/zen3/software/Bowtie2/2.4.5-GCC-11.3.0/bin/bowtie2-align-s |
| R | 4.2.1 |
| step | start | end | elapsed |
|---|---|---|---|
| Read and reference sequence file input | 09:26:30 05 Jul 2024 | 09:27:01 05 Jul 2024 | 31 seconds |
| Read alignment to reference genome | 09:27:01 05 Jul 2024 | 09:28:09 05 Jul 2024 | 1 minute 8 seconds |
| Preprocessing alignments for candidate junction identification | 09:28:09 05 Jul 2024 | 09:28:30 05 Jul 2024 | 21 seconds |
| Preliminary analysis of coverage distribution | 09:28:30 05 Jul 2024 | 09:29:47 05 Jul 2024 | 1 minute 17 seconds |
| Identifying junction candidates | 09:29:47 05 Jul 2024 | 09:29:50 05 Jul 2024 | 3 seconds |
| Re-alignment to junction candidates | 09:29:50 05 Jul 2024 | 09:30:06 05 Jul 2024 | 16 seconds |
| Resolving best read alignments | 09:30:06 05 Jul 2024 | 09:30:39 05 Jul 2024 | 33 seconds |
| Creating BAM files | 09:30:39 05 Jul 2024 | 09:31:43 05 Jul 2024 | 1 minute 4 seconds |
| Tabulating error counts | 09:31:43 05 Jul 2024 | 09:32:27 05 Jul 2024 | 44 seconds |
| Re-calibrating base error rates | 09:32:27 05 Jul 2024 | 09:32:29 05 Jul 2024 | 2 seconds |
| Examining read alignment evidence | 09:32:29 05 Jul 2024 | 09:37:33 05 Jul 2024 | 5 minutes 4 seconds |
| Polymorphism statistics | 09:37:33 05 Jul 2024 | 09:37:33 05 Jul 2024 | 0 seconds |
| Output | 09:37:33 05 Jul 2024 | 09:37:41 05 Jul 2024 | 8 seconds |
| Output :: Mutation Prediction | 09:37:33 05 Jul 2024 | 09:37:33 05 Jul 2024 | 0 seconds |
| Output :: Mutation Annotation | 09:37:33 05 Jul 2024 | 09:37:33 05 Jul 2024 | 0 seconds |
| Total | 11 minutes 11 seconds | ||